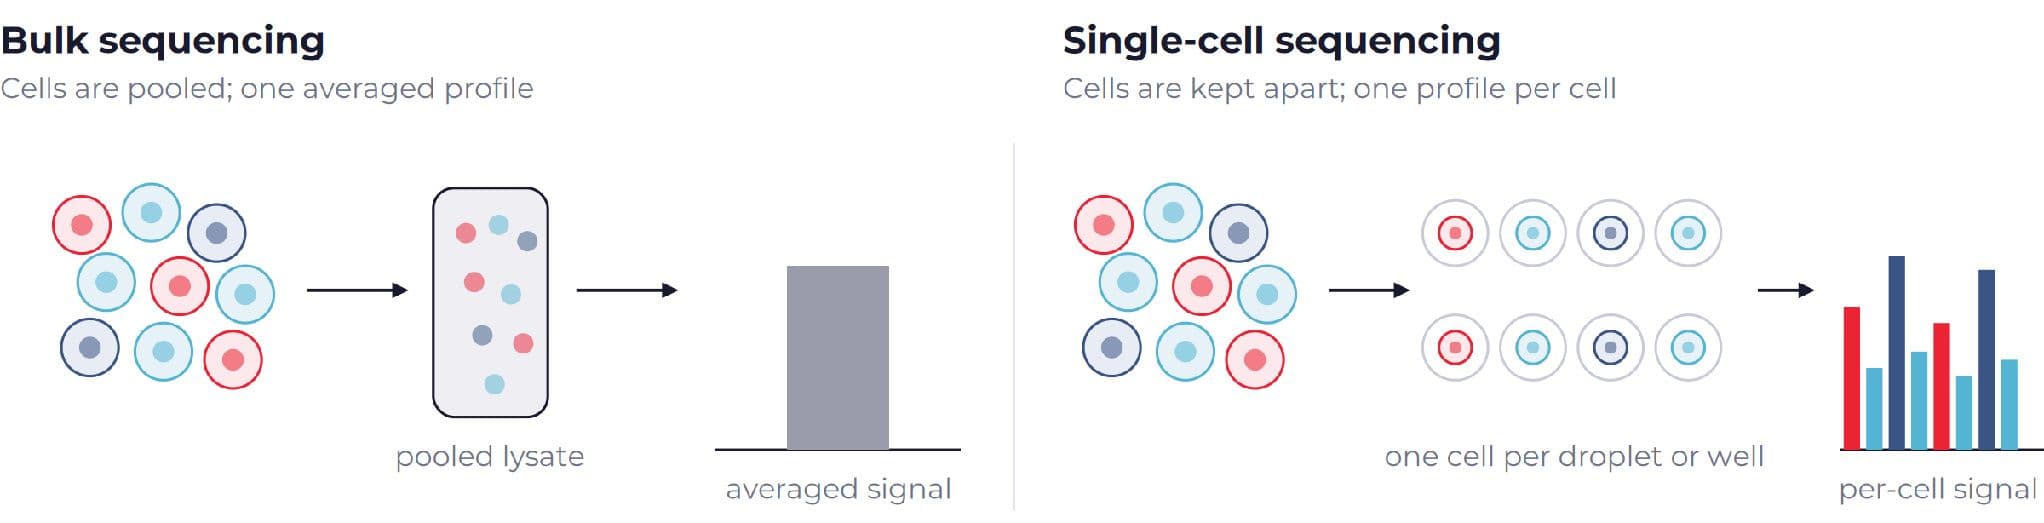
Bulk sequencing averages a population into one number. Single-cell methods keep the differences between cells.
Bulk methods return one averaged measurement across millions of pooled cells, so whatever heterogeneity existed in the sample is collapsed into a single value. Single-cell analysis isolates and reads out individual cells so that those differences survive the experiment, supporting work in tumor clonal evolution, lineage mapping, immune-cell atlasing, measurable residual disease (MRD), cell and gene therapy quality control, and drug-response profiling.
Bulk Sequencing vs Single-Cell Sequencing
Each analyte class shares one constraint: cellular throughput trades against per-cell completeness.
Analyte Class | Molecule measured | Typical throughput | Key limitation |
|---|---|---|---|
Genome | Genomic DNA (mutations, CNV, insertions, deletions) | 10s–10,000s of cells (method dependent) | Whole-genome amplification bottleneck; sharp coverage/throughput trade-off |
Epigenome | Open chromatin, DNA methylation | 1,000s of cells | Very sparse signal per cell |
Transcriptome | mRNA transcripts | 1,000–10,000+ cells per run | Dropout: few mRNA copies per cell |
Proteome | Surface or intracellular protein | 1,000s of cells; tens–hundreds of proteins | Only tens–hundreds of proteins accessible |
Spatial | Any analyte above, plus tissue position | 100s–1,000s of cells per tissue section | Panel size and resolution trade off against area covered |
Multiomics | Two or more analytes from the same cell | 100s–1,000s of cells | Added engineering complexity; lower throughput than single-layer methods |
Correlating separate populations
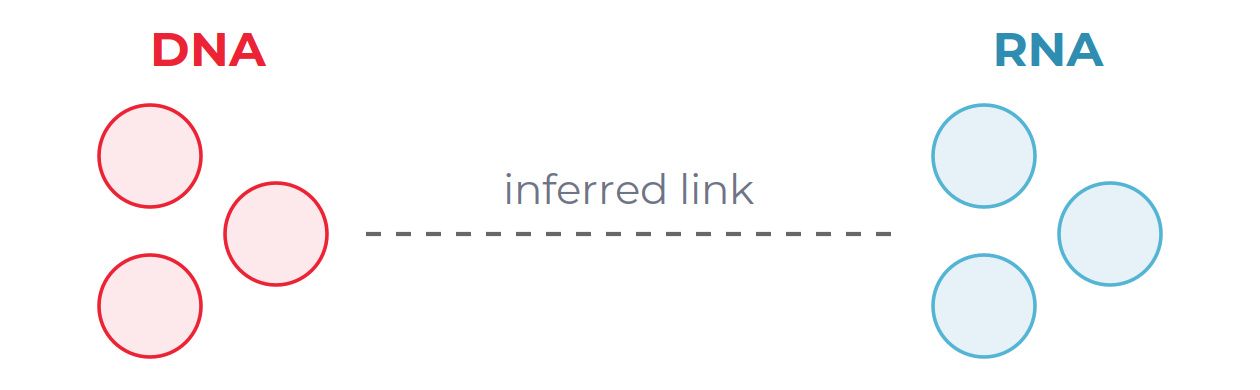
DNA and RNA are measured in different cells from the same sample, so the link between them is inferred, not confirmed.
Measuring the same cell directly
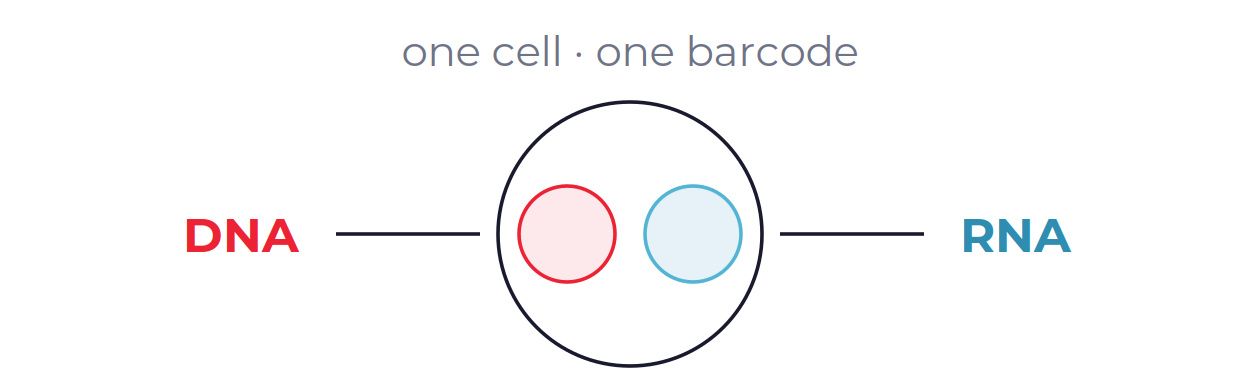
DNA and RNA are measured from the identical single cell, so the link is established directly, at the cost of assay complexity.
Start from the molecular analyte that answers the question, then weigh throughput against completeness.
Three decisions narrow the field: which analyte or analytes matter most, whether throughput or per-cell completeness matters more, and whether two or more analytes, or spatial context, must be linked in the same cell.
The throughput-completeness trade-off · where method families sit
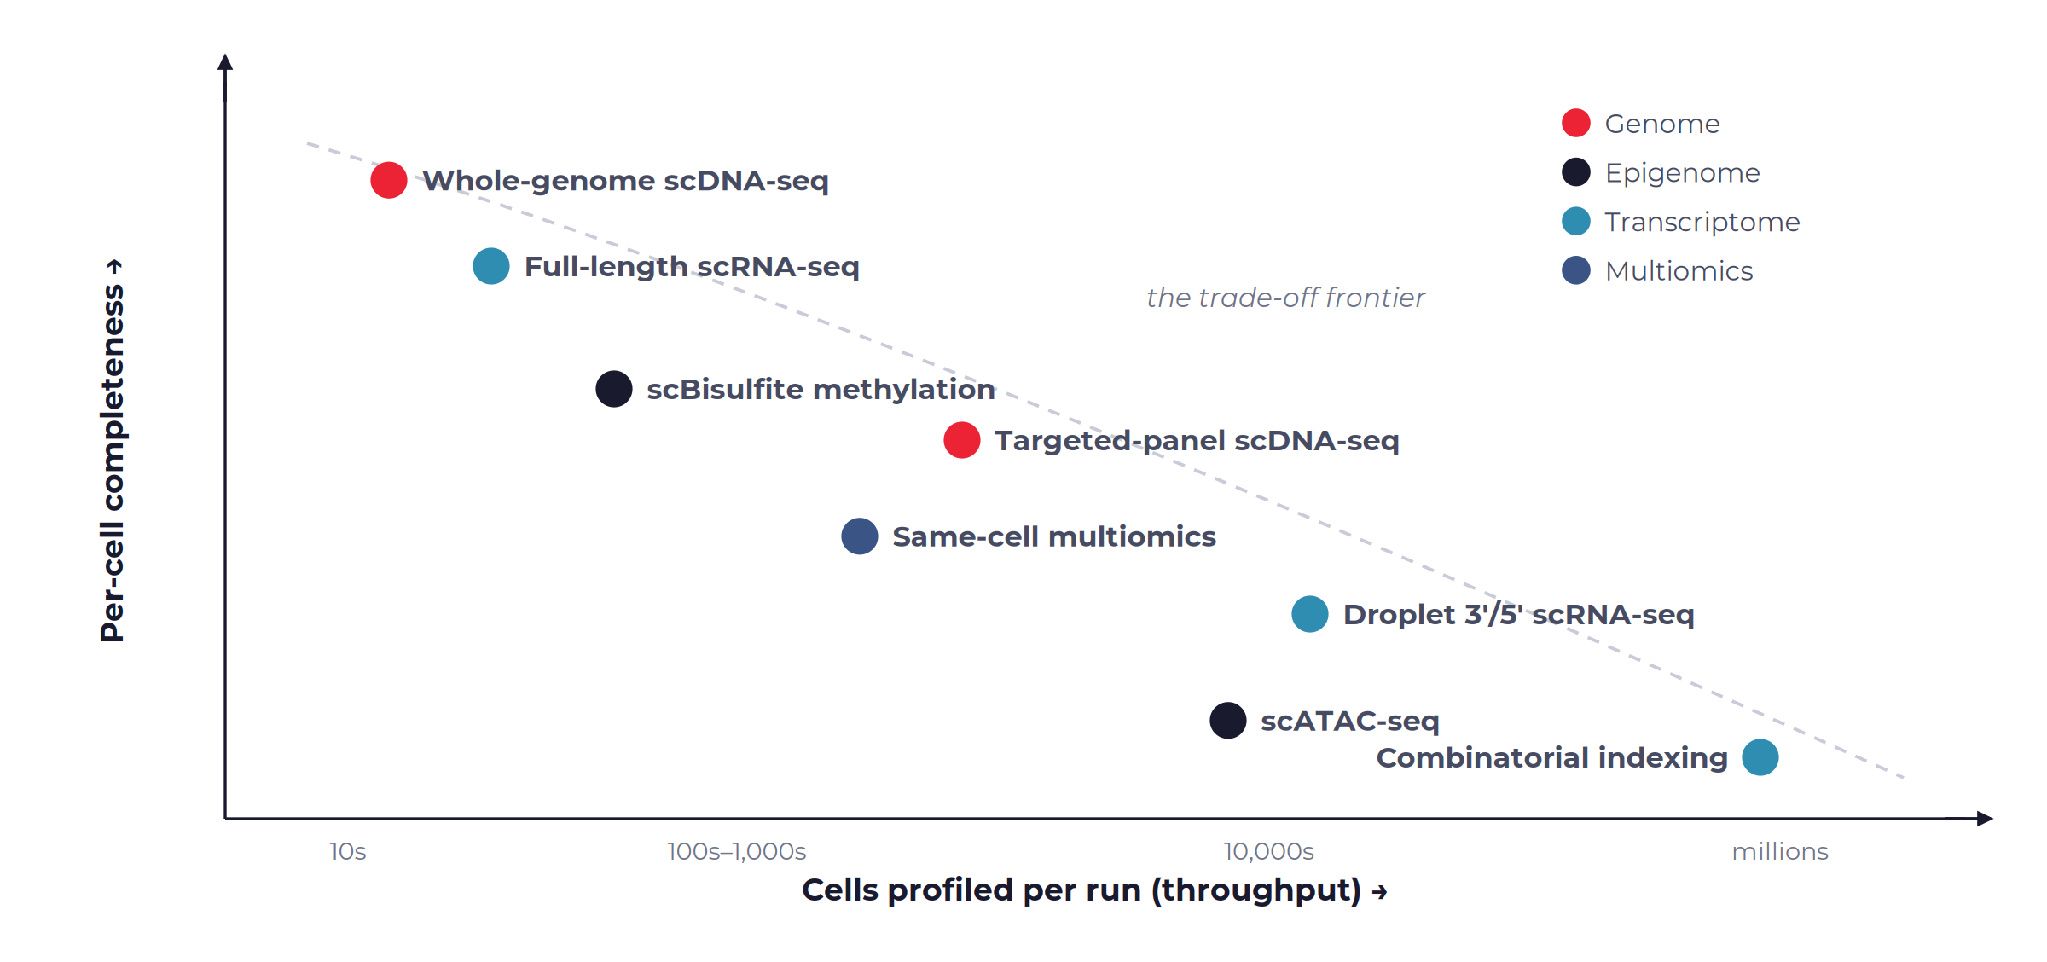
Schematic only: positions are qualitative and summarize the typical published range for each method family. Moving right gains cells per run; moving up gains completeness per cell. Few methods escape the frontier.
What to look for: thousands of cells per run favors droplet, microwell, combinatorial, or targeted-panel methods; complete per-cell coverage (whole genome, whole methylome, whole transcriptome) favors plate-based, full-length, or unbiased methods on fewer cells.